Be sure to fully update your current OS:
- sudo apt-get update
- sudo apt-get upgrade
Fix any broken problems, usually by removing them and re-installing them.
- sudo shutdown -r now
- Then, sudo do-release-upgrade
| Molecular Modeling and Drug Design |
|
|
You can jump between Ubuntu LTS versions but if you are not on the LTS you 'd have to first update to the next Ubuntu version.
Be sure to fully update your current OS:
Fix any broken problems, usually by removing them and re-installing them.
0 Comments
Textbook for Teaching Scientific Computing to Chemistry Students with Python and Jupyter Notebooks7/18/2021 This book is written to teach the subject using Python, Jupyter notebooks, and the SciPy stack which are all open source and cross-platform software; the text assumes no previous computer programming background from the students. The textbook is an open educational resource released under the CC BY-NC-SA 4.0 Creative Commons license and can be freely downloaded at https://github.com/weisscharlesj/SciCompforChemists together with related material.
The article below provides descriptions and philosophy behind the structure and content of the book along with insights and advice based on its use in an undergraduate course: https://pubs.acs.org/doi/10.1021/acs.jchemed.0c01071 To compute the MSD with error using GROMACS use the gmx analyze command:
http://manual.gromacs.org/documentation/5.1/onlinehelp/gmx-analyze.html For error bars in a single trajectory (block averaging) use the -ee option: "Option -ee produces error estimates using block averaging. A set is divided in a number of blocks and averages are calculated for each block. The error for the total average is calculated from the variance between averages of the m blocks B_i as follows: error^2 = sum (B_i - <B>)^2 / (m*(m-1)). These errors are plotted as a function of the block size." The command line is: gmx analyze -f msd_all_DPPC.xvg -ee err_dppc Error bars can also be added with the option -errbar, but only if you have multiple replicas of the same system. The errorbars can represent the standard deviation. The command line is: gmx analyze -f msd_all_DPPC.xvg -ee err_dppc -errbar stddev Sofia Have you ever faced the situation where you perform a long-running task on a remote machine, and suddenly your connection drops, the SSH session is terminated, and your work is lost. Well, it has happened to all of us at some point, hasn’t it? Luckily, there is a utility called screen that allows us to resume the sessions.
Screen or GNU Screen is a terminal multiplexer. In other words, it means that you can start a screen session and then open any number of windows (virtual terminals) inside that session. Processes running in Screen will continue to run when their window is not visible even if you get disconnected. The screen package is pre-installed on most Linux distros nowadays. More information and how to use it is here: https://linuxize.com/post/how-to-use-linux-screen/ Alexis Hello all,
Using the app: https://termius.com/ you can connect with ssh to any linux machine (it's a similar app to e.g. Putty in a Windows PC) . So you can connect to your lab desktop and through it you can ssh to a supercomputer or any server you are using to submit/monitor jobs and run any terminal commands you need. Sofia The following tutorial is designed to help you learn how to become a journal reviewer. It is highly recommended as you 'll also learn how reviewers think, so you can better prepare your own manuscripts. It only takes 2-3 hours and it's free.
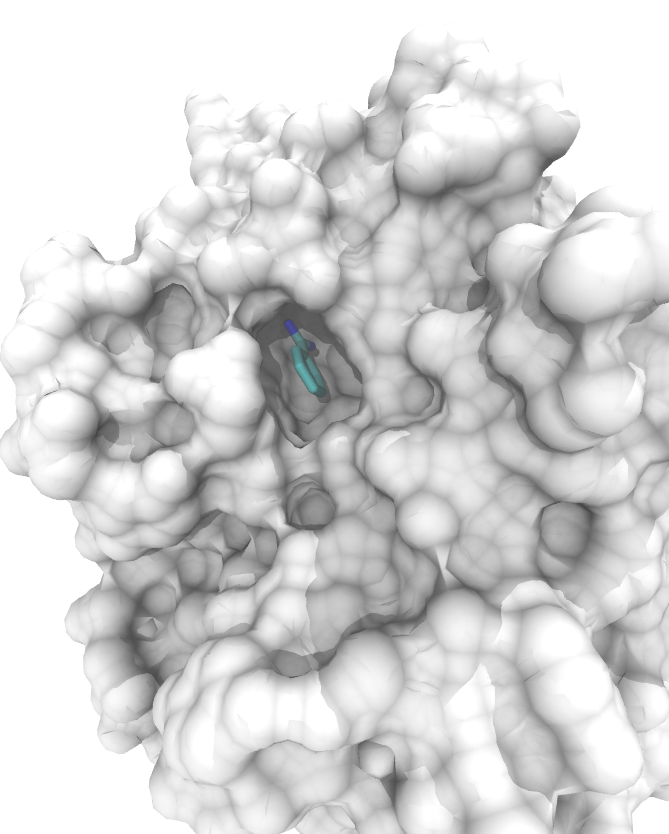
As a Bonus, a certificate is issued if you answer correctly an easy assessment at the end of the tutorial! https://www.acsreviewerlab.org/ Alexis For nice images of proteins in VMD we recommend that you use the Graphics -> Representations -> Drawing Method -> Surf or MSMS visualisation, and then use the "Display -> Display Settings -> Ray Tracing Options -> Amb. Occl -> on" feature. Here you can download MSMS: http://mgltools.scripps.edu/downloads#msms To render: # smaller render, no normalization, high aasamling [tachyonpath] -res 1920 1080 -aasamples 36 -add_skylight 1.0 %s -format PSD48 -o %s.psd # huge render, high aasamling [tachyonpath] -res 12460 9620 -aasamples 36 -add_skylight 1.0 %s -normalize -format PSD48 -o %s.psd # below command can be used to set resolution when you don't want to force a resolution in tachyon display resize 1280 1024 Join Christopher Lipinski and Barry Bunin for a discussion on the "Lipinski Rules of 5," which has become the benchmark by which almost all drug discovery analyses are now compared.
Register here: https://info.collaborativedrug.com/lipinski-bunin-ruleof5-0-0 Experts will discuss:
Very often protein crystallography results in ambiguous regions of electron density.
To check that your protein structure is adequately represented by electron dements maps please see the following tutorial in pymol. Zoe In order to create high quality figures with VMD, go to: File > Render and use the Tachyon module in the File Render Controls panel.
Also, in order to increase the resolution of the figure, go to Render Command, in the same panel, and add the flag: -res xxx yyy, defining the number of pixels of the image. Example: "/usr/local/lib/vmd/tachyon_LINUXAMD64" -aasamples 12 %s -format TARGA -res 1600 1200 -o %s.tga Christina |
Alexis, Michalis, Saman, Nishita, Filippo, Giuseppe, Zoelab group members! Click to set custom HTML
Archives
August 2026
Categories
All
|